InVivoMAb anti-mouse/human CD11b
ILC2 cell purification
Flow cytometry
Product Description
Specifications
| Isotype | Rat IgG2b, κ |
|---|---|
| Recommended Isotype Control(s) | InVivoMAb rat IgG2b isotype control, anti-keyhole limpet hemocyanin |
| Recommended Dilution Buffer | InVivoPure pH 7.0 Dilution Buffer |
| Conjugation | This product is unconjugated. Conjugation is available via our Antibody Conjugation Services. |
| Immunogen | B10 mouse spleen cells enriched for T cells |
| Reported Applications |
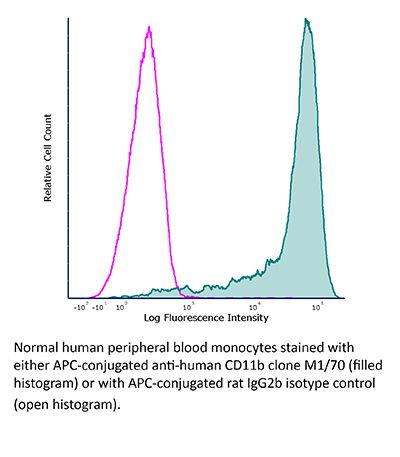
in vivo CD11b neutralization ILC2 cell purification Flow cytometry |
| Formulation |
PBS, pH 7.0 Contains no stabilizers or preservatives |
| Endotoxin |
≤1EU/mg (≤0.001EU/μg) Determined by LAL assay |
| Purity |
≥95% Determined by SDS-PAGE |
| Sterility | 0.2 µm filtration |
| Production | Purified from cell culture supernatant in an animal-free facility |
| Purification | Protein G |
| RRID | AB_1107582 |
| Molecular Weight | 150 kDa |
| Storage | The antibody solution should be stored at the stock concentration at 4°C. Do not freeze. |
| Need a Custom Formulation? | See All Antibody Customization Options |
Application References
-
Li, W., et al (2012). "Intravital 2-photon imaging of leukocyte trafficking in beating heart" J Clin Invest 122(7): 2499-2508.
PubMed
Two-photon intravital microscopy has substantially broadened our understanding of tissue- and organ-specific differences in the regulation of inflammatory responses. However, little is known about the dynamic regulation of leukocyte recruitment into inflamed heart tissue, largely due to technical difficulties inherent in imaging moving tissue. Here, we report a method for imaging beating murine hearts using intravital 2-photon microscopy. Using this method, we visualized neutrophil trafficking at baseline and during inflammation. Ischemia reperfusion injury induced by transplantation or transient coronary artery ligation led to recruitment of neutrophils to the heart, their extravasation from coronary veins, and infiltration of the myocardium where they formed large clusters. Grafting hearts containing mutant ICAM-1, a ligand important for neutrophil recruitment, reduced the crawling velocities of neutrophils within vessels, and markedly inhibited their extravasation. Similar impairment was seen with the inhibition of Mac-1, a receptor for ICAM-1. Blockade of LFA-1, another ICAM-1 receptor, prevented neutrophil adherence to endothelium and extravasation in heart grafts. As inflammatory responses in the heart are of great relevance to public health, this imaging approach holds promise for studying cardiac-specific mechanisms of leukocyte recruitment and identifying novel therapeutic targets for treating heart disease.
-
Liu, B., et al (2015). "Collaborative interactions between type 2 innate lymphoid cells and antigen-specific CD4+ Th2 cells exacerbate murine allergic airway diseases with prominent eosinophilia" J Immunol 194(8): 3583-3593.
PubMed
Type-2 innate lymphoid cells (ILC2s) and the acquired CD4(+) Th2 and Th17 cells contribute to the pathogenesis of experimental asthma; however, their roles in Ag-driven exacerbation of chronic murine allergic airway diseases remain elusive. In this study, we report that repeated intranasal rechallenges with only OVA Ag were sufficient to trigger airway hyperresponsiveness, prominent eosinophilic inflammation, and significantly increased serum OVA-specific IgG1 and IgE in rested mice that previously developed murine allergic airway diseases. The recall response to repeated OVA inoculation preferentially triggered a further increase of lung OVA-specific CD4(+) Th2 cells, whereas CD4(+) Th17 and ILC2 cell numbers remained constant. Furthermore, the acquired CD4(+) Th17 cells in Stat6(-/-)/IL-17-GFP mice, or innate ILC2s in CD4(+) T cell-ablated mice, failed to mount an allergic recall response to OVA Ag. After repeated OVA rechallenge or CD4(+) T cell ablation, the increase or loss of CD4(+) Th2 cells resulted in an enhanced or reduced IL-13 production by lung ILC2s in response to IL-25 and IL-33 stimulation, respectively. In return, ILC2s enhanced Ag-mediated proliferation of cocultured CD4(+) Th2 cells and their cytokine production, and promoted eosinophilic airway inflammation and goblet cell hyperplasia driven by adoptively transferred Ag-specific CD4(+) Th2 cells. Thus, these results suggest that an allergic recall response to recurring Ag exposures preferentially triggers an increase of Ag-specific CD4(+) Th2 cells, which facilitates the collaborative interactions between acquired CD4(+) Th2 cells and innate ILC2s to drive the exacerbation of a murine allergic airway diseases with an eosinophilic phenotype.
-
Becker, A. M., et al (2015). "ADAM17 limits the expression of CSF1R on murine hematopoietic progenitors" Exp Hematol 43(1): 44-52 e41-43.
PubMed
All-lymphoid progenitors (ALPs) yield few myeloid cells in vivo, but readily generate such cells in vitro. The basis for this difference remains unknown. We hypothesized that ALPs limit responsiveness to in vivo concentrations of myeloid-promoting cytokines by reducing expression of the corresponding receptors, potentially through posttranscriptional mechanisms. Consistent with such a mechanism, ALPs express higher levels of CSF1R transcripts than their upstream precursors, yet show limited cell-surface protein expression of colony-stimulating factor 1 receptor (CSF1R). All-lymphoid progenitors and other hematopoietic progenitors deficient in A disintegrin and metalloproteinase domain 17 (ADAM17), display elevated cell surface CSF1R expression. ADAM17(-/-) ALPs, however, fail to yield myeloid cells upon transplantation into irradiated recipients. Moreover, ADAM17(-/-) ALPs yield fewer macrophages in vitro than control ALPs at high concentrations of macrophage colony stimulating factor. Mice with hematopoietic-specific deletion of ADAM17 have normal numbers of myeloid and lymphoid progenitors and mature cells in vivo. These data demonstrate that ADAM17 limits CSF1R protein expression on hematopoietic progenitors, but that compensatory mechanisms prevent elevated CSF1R levels from altering lymphoid progenitor potential.
-
Yokota, N., et al (2014). "Contributions of thrombin targets to tissue factor-dependent metastasis in hyperthrombotic mice" J Thromb Haemost 12(1): 71-81.
PubMed
BACKGROUND: Tumor cell tissue factor (TF)-initiated coagulation supports hematogenous metastasis by fibrin formation, platelet activation and monocyte/macrophage recruitment. Recent studies identified host anticoagulant mechanisms as a major impediment to successful hematogenous tumor cell metastasis. OBJECTIVE: Here we address mechanisms that contribute to enhanced metastasis in hyperthrombotic mice with functional thrombomodulin deficiency (TM(Pro) mice). METHODS: Pharmacological and genetic approaches were combined to characterize relevant thrombin targets in a mouse model of experimental hematogenous metastasis. RESULTS: TF-dependent, but contact pathway-independent, syngeneic breast cancer metastasis was associated with marked platelet hyperreactivity and formation of leukocyte-platelet aggregates in immune-competent TM(Pro) mice. Blockade of CD11b or genetic deletion of platelet glycoprotein Ibalpha excluded contributions of these receptors to enhanced platelet-dependent metastasis in hyperthrombotic mice. Mice with very low levels of the endothelial protein C receptor (EPCR) did not phenocopy the enhanced metastasis seen in TM(Pro) mice. Genetic deletion of the thrombin receptor PAR1 or endothelial thrombin signaling targets alone did not diminish enhanced metastasis in TM(Pro) mice. Combined deficiency of PAR1 on tumor cells and the host reduced metastasis in TM(Pro) mice. CONCLUSIONS: Metastasis in the hyperthrombotic TM(Pro) mouse model is mediated by platelet hyperreactivity and contributions of PAR1 signaling on tumor and host cells.
Product Citations
-
Spatial dynamics of viral replication drive biphasic lymphoid activation and systemic alphavirus immune responses.
In PLoS Pathog on 12 August 2026 by Jacoby, M. L., Sun, C., et al.
PubMed
Systemic type I interferon (IFN-I) and pro-inflammatory cytokine responses are critical for limiting mosquito-borne RNA virus viral replication and disease, yet mechanisms initiating these responses and their linkage to viral tropism for lymphoid tissues are poorly understood. Here, we reveal that virus-cell interactions and specific host signaling pathways in lymphoid tissues are determinants of systemic cytokine induction. Using Venezuelan equine encephalitis virus (VEEV) mutants with defined lymph node (LN) tropism in mice, we show that VEEV LN infection positively associates with systemic IFN-I and other proinflammatory cytokine induction. Maximal responses require coordinated activation of multiple IRF7-dependent Toll-like receptor and RIG-I-like receptor pathways. Combining targeted cell depletion and Visium HD spatial transcriptomics viral gene quantitation, we uncover a spatiotemporal, biphasic innate immune response to VEEV infection, with initial cytokine induction driven by VEEV replication in dendritic cell and monocyte/macrophage subtypes within the LN subcapsular sinus. Surprisingly, active virus replication is curtailed in this LN compartment before 16hpi, when cytokine induction becomes dominated by non-VEEV-replicating bystander cells including plasmacytoid dendritic cells. These findings have direct implications for intervention against viral diseases characterized by lymphoid infection and subsequent spread to sites of terminal disease.
-
TLR7 Inhibition Limits Ischemic Cardiac Injury by Disrupting ITGAM-Dependent Immune-Endothelial Interactions.
In JACC Basic Transl Sci on 1 August 2026 by Li, Y., Yang, Y., et al.
PubMed
Percutaneous coronary intervention, such as stenting, restores coronary perfusion after myocardial ischemia but also induces ischemia-reperfusion (I/R) injury, in part driven by innate immune activation. Here, we identified Toll-like receptor 7 (TLR7), an endosomal sensor of single-stranded RNA, as a key mediator of postischemic inflammation. Genetic deletion or pharmacologic inhibition of TLR7 with enpatoran reduced inflammation and myocardial infarction (MI) size and improved cardiac function 24 hours after I/R in mice. Single-nucleus RNA sequencing demonstrated coordinated induction of TLR7-responsive inflammatory endothelial and myeloid cell programs, including increased endothelial ITGAM expression and leukocyte adhesion. TLR7/8 activation promoted ITGAM-dependent endothelial-leukocyte interactions under shear stress, whereas ITGAM blockade reduced immune cell infiltration and limited MI injury. These findings identify the TLR7-ITGAM signaling axis as a central driver of endothelial-leukocyte crosstalk in myocardial I/R injury and support TLR7 inhibition as a promising therapeutic strategy to mitigate ischemic MI after percutaneous coronary intervention.
-
Immobilized IgG-containing immune complexes require platelets to recruit neutrophils during inflammation.
In J Clin Invest on 2 February 2026 by Bellio, M., Allaeys, I., et al.
PubMed
During vascular injury, platelets are essential for halting bleeding and recruiting neutrophils to prevent microbial invasion. However, in antibody-mediated autoimmune diseases occurring without vascular damage, neutrophils infiltrate tissues and contribute to pathology. Here, we investigated whether the dependence of neutrophils on platelets is conserved in the context of antibody-driven inflammation. Using human cells from individuals with rheumatoid arthritis and a microfluidic system mimicking physiological shear over IgG-containing immune complexes, we demonstrate that despite expressing Fc receptors, neutrophils required platelets to stably adhere to immune complexes under flow. Platelet Fcγ receptor 2a (FcγRIIA) binding was critical for resisting shear stress, while neutrophils used FcγRIIA and FcγRIIIB for immune complex recognition. Platelet P-selectin binding to neutrophil P-selectin glycoprotein ligand 1 (PSGL-1) was essential for recruitment, whereas macrophage-1 antigen (Mac-1) was dispensable. In a mouse model of autoantibody-mediated arthritis, intravital imaging confirmed that neutrophil recruitment relied on PSGL-1. Importantly, expression of FcγRIIA aggravated arthritis, and blockade of PSGL-1, but not Mac-1, in these mice abrogated both the platelet and neutrophil interactions and disease. These findings identify key molecular interactions in platelet-neutrophil cooperation and reveal that platelets are essential enablers of FcR-mediated neutrophil adhesion in antibody-driven inflammation.
-
Mac-1 Promotes Neutrophil Extracellular Traps Formation via ERK Phosphorylation in Renal Ischemia-Reperfusion Injury.
In Inflammation on 1 December 2025 by He, L., Wang, J., et al.
PubMed
Neutrophils are key effector cells in the pathogenesis of acute kidney injury (AKI), where neutrophil extracellular traps (NETs) play a critical role. Mac-1 (CD11b/CD18), a leukocyte surface integrin, is known to mediate inflammation. We hypothesized that Mac-1 regulates NETs formation and exacerbates ischemic AKI. The expression of Mac-1 and NETs was analyzed in peripheral blood and renal tissues from patients with kidney diseases. Mac-1 knockout (Mac-1⁻/⁻) mice were used to establish a bilateral renal ischemia-reperfusion injury (IRI) model and compared with wild-type (WT) controls. Primary neutrophils were isolated from bone marrow in vitro to investigate the mechanisms underlying NETs formation. CD11b and NETs were both implicated in patients with either AKI following cardiac surgery or interstitial nephritis. In a murine bilateral renal IRI model, NETs formation was detected in WT kidneys at 24 h, and DNase I treatment significantly alleviated renal injury. Compared to WT mice, Mac-1⁻/⁻ mice exhibited reduced dysfunction, inflammation, and NETs infiltration. In vitro, Mac-1⁻/⁻ neutrophils showed decreased PMA-induced NETs formation and reduced ERK phosphorylation. Furthermore, treatment with anti-Mac-1 antibody (M1/70) in the murine IRI model significantly attenuated kidney inflammation and injury. Mac-1 exacerbates kidney inflammation and dysfunction in ischemic AKI by mediating NETs formation through an ERK phosphorylation-dependent mechanism. Blockade of Mac-1 using the antibody M1/70 effectively ameliorates this injury, highlighting its potential as a therapeutic target.